Export and convenience functions for ancestral reconstructions
Source:R/ancestral_generics.R
write.ancestral.Rdwrite.ancestral allows to export ancestral reconstructions. It writes
out the tree, a tab delimited text file with the probabilities, an alignment
of the the extant sequences and the most likely ancestral states. And a
unaligned fasta file containg the most likely ancestral states.
ancestral generates an object of class ancestral.
Usage
write.ancestral(x, file = "ancestral")

as.ancestral(tree, prob, align, ...)

# S3 method for class 'ancestral'
print(x, ...)

# S3 method for class 'ancestral'
as.phyDat(x, ...)

# S3 method for class 'ancestral'
as.data.frame(x, ..., terminal = FALSE)
Arguments
- x
an object of class ancestral
- file
a file name. File endings are added.
- tree
an object of class phylo.
- prob
an data.frame containing a matrix of posterior probabilities for each state and site.
- align
an object of class phyDat.
- ...
Further arguments passed to or from other methods.
- terminal
logical, should terminal nodes included.
Details
This allows also to read in reconstruction made by iqtree to use the plotting capabilities of R.
Examples
data(Laurasiatherian)
fit <- pml_bb(Laurasiatherian[,1:100], "JC", rearrangement = "none")
anc_ml <- anc_pml(fit)
write.ancestral(anc_ml)
# Can be also results from iqtree
align <- read.phyDat("ancestral_all.fasta", format="fasta")
tree <- read.tree("ancestral_tree.nwk")
df <- read.table("ancestral_state.tsv.gz", header=TRUE)
anc_ml_disc <- as.ancestral(tree, df, align)
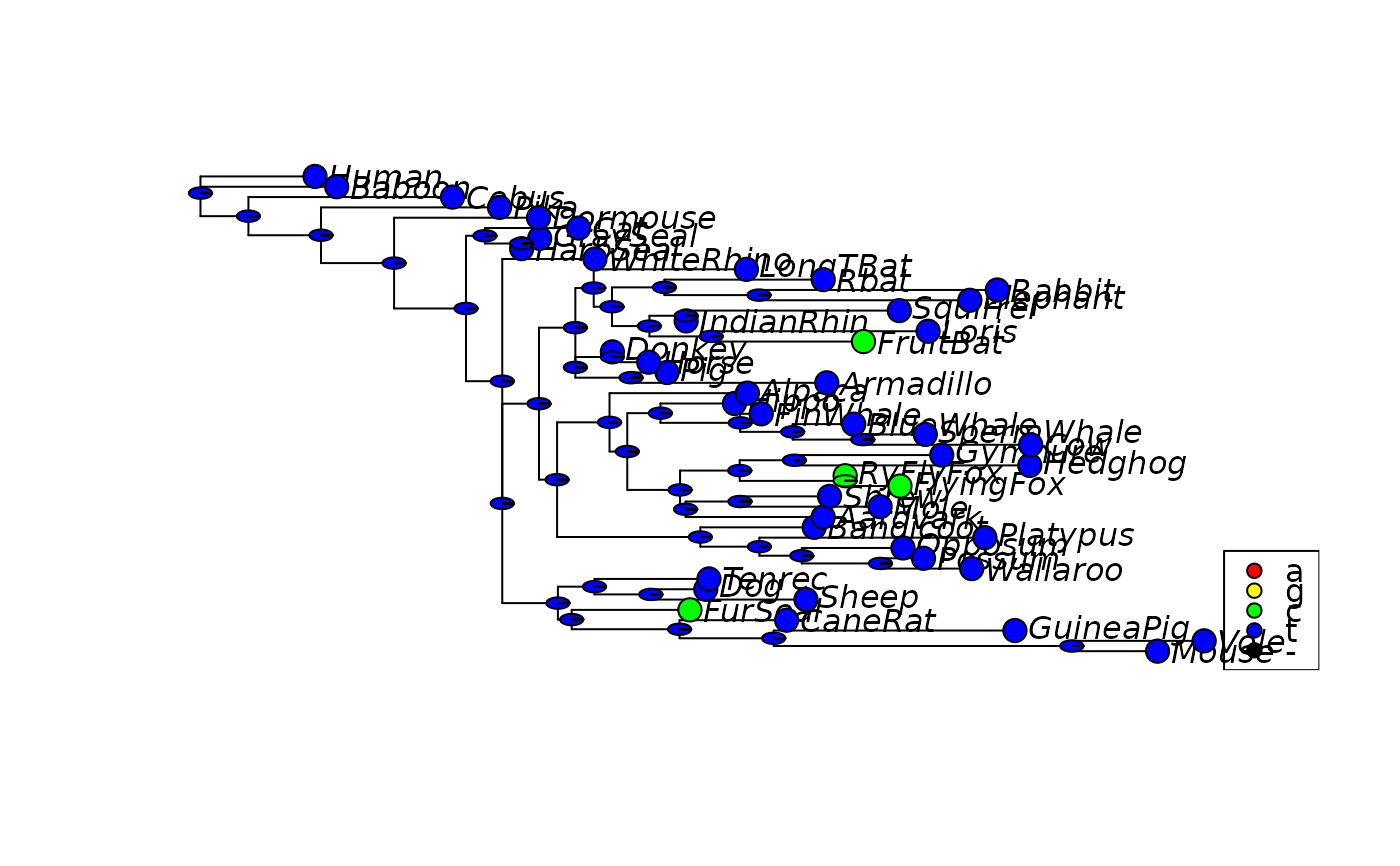
plotAnc(anc_ml_disc, 20)
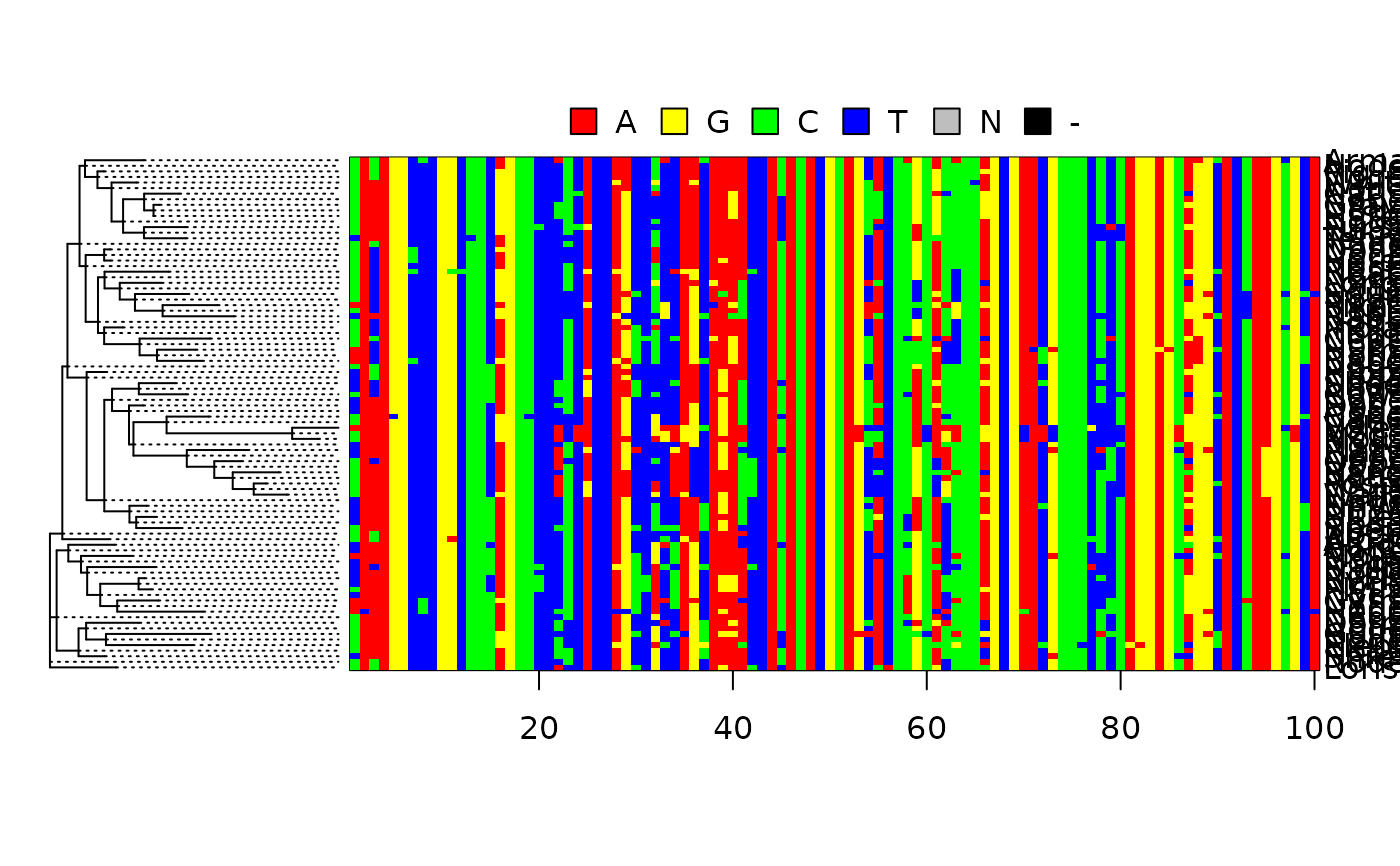
 heatmapp(tree, align, nodes=TRUE)
 unlink(c("ancestral_all.fasta", "ancestral_tree.nwk",
"ancestral_state.tsv.gz", "ancestral_state.fasta"))